Twisted Ethylene with MRSF-TDDFT
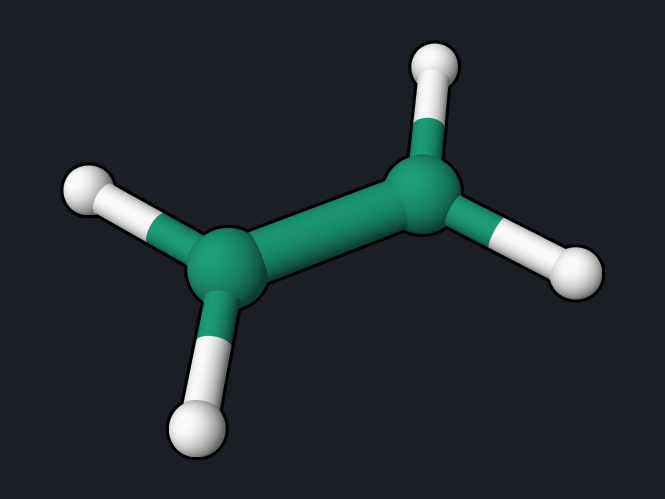
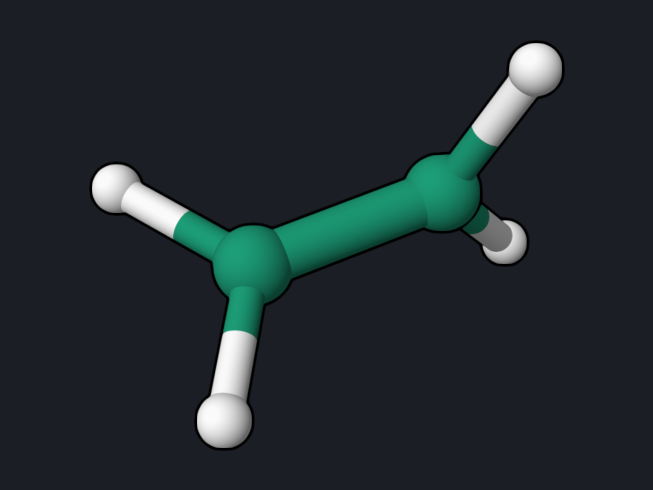
Twisting ethylene about its C=C bond breaks the π bond. At 90° the π and π* orbitals have the same energy and the ground state becomes a diradical. A method built on one closed-shell determinant cannot describe this ground state. MRSF-TDDFT obtains the ground and excited states together from a high-spin triplet reference, so the same input works at every twist angle.
Copy and run
Planar ethylene at its PBE0/6-31G* geometry:
mrsf(nstate=3)/bhhlyp/6-31g*
geom="""
C      0.000000     0.000000     0.664311
C      0.000000     0.000000    -0.664311
H      0.000000     0.923944     1.237411
H      0.000000    -0.923944     1.237411
H      0.000000     0.923944    -1.237411
H      0.000000    -0.923944    -1.237411
"""
The same molecule with one CH₂ group rotated by 90°:
mrsf(nstate=3)/bhhlyp/6-31g*
geom="""
C      0.000000     0.000000     0.664311
C      0.000000     0.000000    -0.664311
H      0.000000     0.923944     1.237411
H      0.000000    -0.923944     1.237411
H      0.923944     0.000000    -1.237411
H     -0.923944     0.000000    -1.237411
"""
openqp ethylene-planar.oqp
openqp ethylene-twisted.oqp
mrsf(nstate=3) requests three singlet states, S0, S1, and S2. BHHLYP is the
functional most commonly used with MRSF-TDDFT. OpenQP builds
the required high-spin ROHF triplet reference automatically. Each run takes
about 3 seconds.
What you get
The ENERGIES AND STATES block of each log lists the states. For planar
ethylene:
   PyOQP triplet ROHF reference (internal)  -78.3689760267
   PyOQP S0                                 -78.5257616296
   PyOQP S1                                 -78.2178196463
   PyOQP S2                                 -78.2165027669
and for twisted ethylene:
   PyOQP triplet ROHF reference (internal)  -78.4201554151
   PyOQP S0                                 -78.3957310053
   PyOQP S1                                 -78.3138521561
   PyOQP S2                                 -78.3115552147
The first line is the triplet reference used internally to build the response states; it is not one of the computed singlet states. Converted to energy differences:
| Quantity | Planar (0°) | Twisted (90°) |
|---|---|---|
| S0 → S1 vertical excitation | 8.38 eV | 2.23 eV |
| S0 → S2 vertical excitation | 8.42 eV | 2.29 eV |
| S0 energy relative to planar S0 | 0 | 3.54 eV |
The twist curve
Repeating the calculation every 15° gives the potential-energy curves below. The bond lengths and H–C–H angles are kept fixed, so this is a rigid twist; the relaxed barrier is lower.
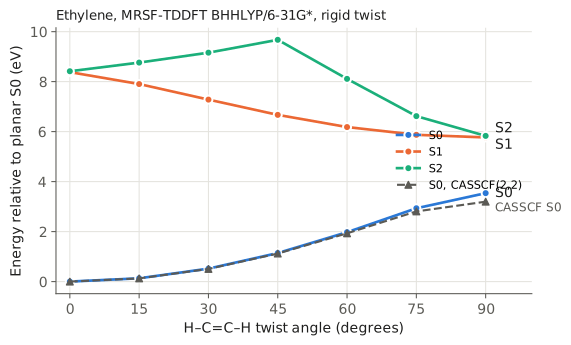
The dashed curve is the S0 energy from CASSCF(2,2) with the same basis set at the same geometries, with two electrons in the π and π* orbitals. At 90° it lies 3.20 eV above its planar value, compared with 3.54 eV for MRSF-TDDFT S0; the two methods agree on the shape of the curve, while CASSCF(2,2) lacks the dynamic correlation that MRSF-TDDFT includes through the functional. The CASSCF input for 90° is
casscf/6-31g* cas(active_electrons=2,active_orbitals=2)
geom="""
C      0.000000     0.000000     0.664311
C      0.000000     0.000000    -0.664311
H      0.000000     0.923944     1.237411
H      0.000000    -0.923944     1.237411
H     -0.923944     0.000000    -1.237411
H      0.923944     0.000000    -1.237411
"""
Download ethylene-twisted-casscf.oqp; for the other angles, use the scan script below with this first line.
| Twist | S0 (eV) | S1 (eV) | S2 (eV) |
|---|---|---|---|
| 0° | 0.000 | 8.380 | 8.415 |
| 15° | 0.132 | 7.903 | 8.762 |
| 30° | 0.518 | 7.280 | 9.159 |
| 45° | 1.142 | 6.670 | 9.673 |
| 60° | 1.976 | 6.182 | 8.110 |
| 75° | 2.933 | 5.872 | 6.619 |
| 90° | 3.538 | 5.766 | 5.829 |
All energies are relative to S0 of planar ethylene. S0 rises as the π bond breaks, while S1 falls, and the S0–S1 gap closes from 8.4 eV to 2.2 eV. This smooth, continuous description of S0 near a broken bond is what MRSF-TDDFT is used for in photochemistry, for example in searches for conical intersections (see BaekA Multistate MECI). The S2 curve changes character between 45° and 75°; with the 6-31G* basis set, the upper states mix valence and Rydberg-like character and should be interpreted with care.
To make the scan inputs, this Python script writes one .oqp file per angle by
rotating the two hydrogen atoms on the second carbon about the C=C axis:
import math

base = [("C", 0.0, 0.0, 0.664311), ("C", 0.0, 0.0, -0.664311),
        ("H", 0.0, 0.923944, 1.237411), ("H", 0.0, -0.923944, 1.237411),
        ("H", 0.0, 0.923944, -1.237411), ("H", 0.0, -0.923944, -1.237411)]

for deg in range(0, 91, 15):
    t = math.radians(deg)
    lines = []
    for i, (el, x, y, z) in enumerate(base):
        if i >= 4:  # hydrogens on the second carbon
            x, y = x * math.cos(t) - y * math.sin(t), x * math.sin(t) + y * math.cos(t)
        lines.append(f"{el} {x:12.6f} {y:12.6f} {z:12.6f}")
    with open(f"twist{deg:02d}.oqp", "w") as f:
        f.write("mrsf(nstate=3)/bhhlyp/6-31g*\ngeom=\"\"\"\n" + "\n".join(lines) + "\n\"\"\"\n")
for f in twist*.oqp; do openqp "$f"; done
Try next
- Add
energy(T0)to compute the triplet states,mrsf(nstate=3)/bhhlyp/6-31g* energy(T0). For the twisted input, T0 is −78.3961121 hartree, 0.010 eV below S0, as expected for a diradical. - Replace the first line with
tddft(nstate=3)/bhhlyp/6-31g*to use ordinary TDDFT from the closed-shell ground state. At 0° it works; at 90° the closed-shell reference is no longer a good description of S0, and in our test run the lowest root fell to 0.43 eV and the Davidson solver stopped without converging.
See also: MRSF-TDDFT, Spin-orbit coupling example.