Formaldehyde: Excited-State Geometry, Absorption, and Emission
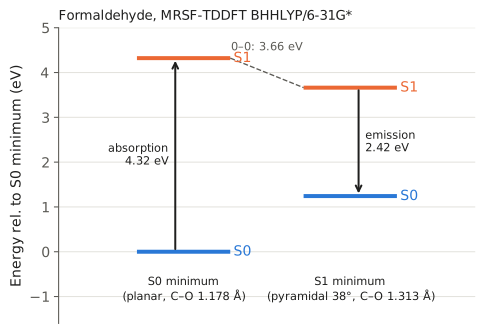
Optimizing a molecule on an excited state gives its relaxed excited-state structure and, from that structure, the emission energy. Together with the ground-state minimum this yields the three numbers that describe a band in absorption and fluorescence spectra: the vertical absorption, the vertical emission, and the adiabatic (0–0) energy. This example does both optimizations for formaldehyde with MRSF-TDDFT.
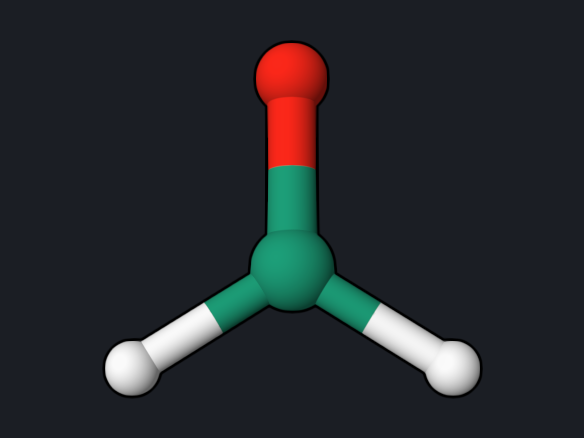
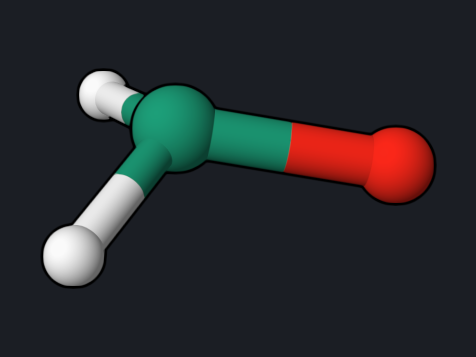
Step 1: optimize the ground state
mrsf(nstate=3)/bhhlyp/6-31g* opt(S0)
geom="""
C      0.000000     0.000000    -0.526342
O      0.000000     0.000000     0.676110
H      0.000000     0.937620    -1.119884
H      0.000000    -0.937620    -1.119884
"""
Download formaldehyde-s0-mrsf-opt.oqp
opt(S0) optimizes the MRSF-TDDFT ground state, so both optimizations use
the same method. The run converges in 6 steps (about 7 seconds).
Step 2: optimize the first excited singlet
mrsf(nstate=3)/bhhlyp/6-31g* opt(S1)
geom="""
C      0.000000     0.000000    -0.526342
O      0.150000     0.000000     0.676110
H      0.000000     0.937620    -1.119884
H      0.000000    -0.937620    -1.119884
"""
Download formaldehyde-s1-opt.oqp
opt(S1) follows the physical first excited singlet, using analytic MRSF-TDDFT
gradients. The oxygen atom starts 0.15 Å out of the plane, so the optimizer is
not held at the planar structure by symmetry; the S1 minimum is pyramidal. The run converges in 10 steps (about
11 seconds):
    9    -114.28784263      -0.00001059       0.003241       0.005782       0.000344       0.000798
   10    -114.28784362      -0.00000100       0.000815       0.001699       0.000058       0.000153
What you get
The ENERGIES AND STATES block at the end of each run gives S0 and S1 at the
optimized geometry:
| Structure | E(S0) (hartree) | E(S1) (hartree) |
|---|---|---|
| S0 minimum | −114.4224437 | −114.2636198 |
| S1 minimum | −114.3767751 | −114.2878436 |
| Quantity | MRSF-TDDFT | Experiment |
|---|---|---|
| Vertical absorption, S1 − S0 at the S0 minimum | 4.32 eV | about 4.1 eV (band maximum) |
| Vertical emission, S1 − S0 at the S1 minimum | 2.42 eV | — |
| Adiabatic 0–0 energy (without zero-point correction) | 3.66 eV | 3.49 eV |
| r(C–O) at S0 / S1 minimum | 1.178 / 1.313 Å | 1.208 / 1.32 Å |
| Out-of-plane angle of O at the S1 minimum | 38° | 34° |
Reading the result
- The n→π* state changes the shape. Moving one electron from the oxygen lone pair into the C=O π* orbital weakens the C–O bond, which lengthens by 0.13 Å, and the molecule becomes pyramidal. The same changes appear in the first femtoseconds of the QM/MM dynamics example.
- Absorption and emission differ by 1.9 eV. The relaxation on S1 lowers S1 by 0.66 eV and raises S0 at that geometry by 1.24 eV, a large Stokes shift for such a small molecule.
- The experimental 0–0 energy includes zero-point vibrational energy; add the difference of the S1 and S0 zero-point energies from Hessian runs for a direct comparison.
Try next
- Use
opt(T0)with the same input to optimize the lowest triplet. - Repeat with TDDFT,
tddft(nstate=3)/pbe0/6-31g* opt(S1), and compare the geometry.
See also: Optimization, MRSF-TDDFT.